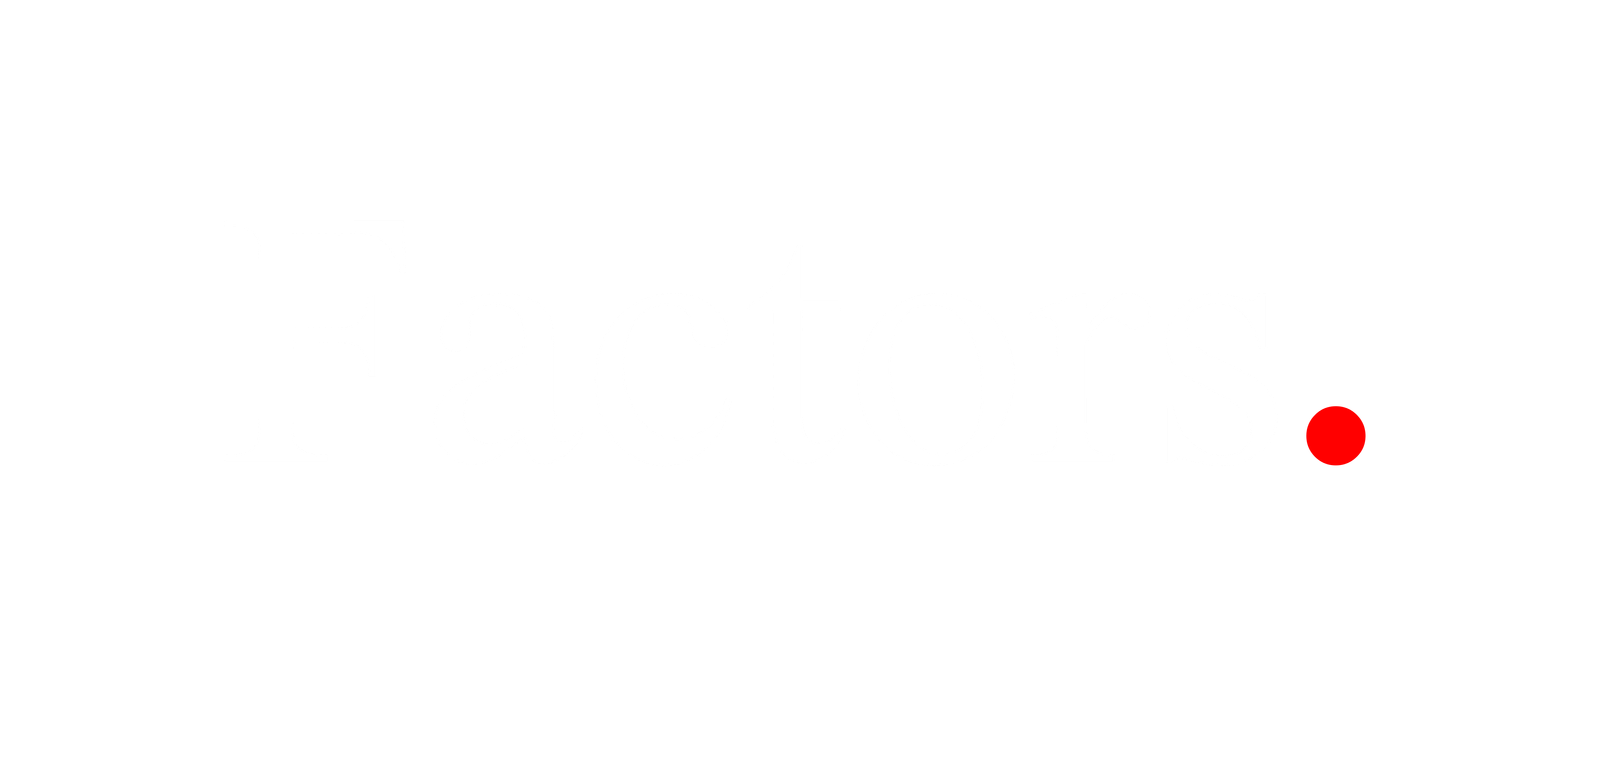
Inside eukaryotic cells, the genome is hierarchically organized into a higher-order chromatin structure rather than existing as unconstrained DNA. DNA wraps around an octameric complex of core histones (H2A, H2B, H3, H4) to form nucleosomes-the fundamental subunits of chromatin. While the hydrophobic octamer core stabilizes the particle, N-terminal histone tails protrude outward into the nuclear environment, exposing them to enzymatic machinery (Figure 1). This exposure allows site-specific post-translational modifications (PTMs)-including acetylation, methylation, phosphorylation, and ubiquitination-to be dynamically deposited by “writers,” removed by “erasers,” and interpreted by “readers.” Within the ‘histone code’ framework, these modifications regulate local chromatin accessibility through distinct modes: lysine acetylation neutralizes positive charges to promote chromatin opening, whereas histone methylation acts in a context-dependent manner to foster either open or condensed chromatin states. Together, these modifications regulate local chromatin spatial accessibility: highly condensed heterochromatin sterically occludes cis-regulatory elements (such as promoters and enhancers), preventing transcription factor binding to enforce gene silencing, whereas localized chromatin remodeling opens euchromatin to permit transcriptional engagement (Figure 1).
Extensive research has identified hundreds of distinct histone PTMs on canonical and variant histones, forming a coordinated network that fine-tunes cell-type-specific gene expression. Crucially, dysregulation of these epigenetic mechanisms drives diverse human pathologies, including cancer, autoimmune conditions, and chronic inflammatory disorders. Because histone PTMs are dynamic and reversible, targeting their regulatory pathways offers a powerful strategy for epigenetic therapy and drug discovery. Global initiatives-such as the ENCODE project and the Roadmap Epigenomics Consortium-have systematically mapped these functional landscapes across diverse cell types and disease states. Building on these foundational maps, elucidating the precise mechanisms governing histone PTM deposition remains essential for decoding gene regulation and identifying novel therapeutic targets.
Out of the vast repertoire of histone PTMs, trimethylation of histone H3 at lysine 4 (H3K4me3) has emerged as one of the most extensively studied marks, featured in over 4,000 PubMed publications to date and widely considered a hallmark of active promoters. Mapping precisely to transcription start sites (TSSs) across eukaryotic evolution, early foundational studies framed H3K4me3 as an instructive driver of gene expression. In this model, H3K4me3 was thought to actively trigger transcription initiation by maintaining open promoter architecture and recruiting chromatin remodelers and basal machinery to assemble RNA Polymerase II (Pol II). However, accumulating evidence challenges this instructive paradigm. Genetic knockout or catalytic inhibition of major MLL/COMPASS family methyltransferases depletes global H3K4me3 without causing a corresponding collapse in steady-state transcription. Likewise, targeted promoter demethylation frequently leaves ongoing transcript synthesis intact. These paradoxical observations create a classic “chicken-and-egg” paradigm: Is H3K4me3 an instructive cause that drives transcription initiation, or a reactive consequence deposited by transcriptional activity?
Resolving this paradox required a model system to overcome a long-standing experimental deadlock. Most chromatin studies rely on constitutively active genes or slow induction paradigms, where transcription and histone modifications change synchronously over extended timeframes, obscuring cause and consequence. Determining whether H3K4me3 deposition precedes, accompanies, or follows transcription demanded high temporal resolution with rapid activation and shutdown. We strategically targeted the inducible human tumor necrosis factor alpha (TNF-α) and interleukin 1 beta (IL-1β) genes, two key inflammatory cytokine genes that exhibit a well-characterized, highly dynamic ‘off-on-off’ expression profile upon lipopolysaccharide (LPS) stimulation, in THP-1 human monocytic cells. Their rapid induction, short mRNA half-lives, and swift resolution provided an ideal experimental window to delineate the precise sequence of molecular events. Evaluating this strategy across two independent genes at their native genomic locations on distinct chromosomes ensured that the observed kinetic trends reflected generalizable biological principles rather than locus-specific artifacts.
Leveraging this temporal framework, our work published in Life Science Alliance (DOI: 10.26508/lsa.202503511) revealed that H3K4me3 accumulates hours after peak transcription rather than coinciding with it, directly challenging models that position this mark as an early requirement for Pol II assembly. Our study provided three distinct lines of evidence supporting this model: First, kinetic analysis revealed clear temporal separation between activation machinery and mark deposition: while NF-κB occupancy, p300 recruitment, histone acetylation (H3K27ac/H4K8ac), and Ser5/Ser2-phosphorylated Pol II correlate peak transcription, MLL1-the methyltransferase responsible for H3K4me3 in these cells-recruitment and H3K4me2/3 accumulation lag significantly behind. Second, we showed that this delayed deposition is strictly transcription-dependent, as pharmacological inhibition of NF-κB (via TPCA-1) or global transcription (via actinomycin D) blunted H3K4me3 accumulation-confirming that active elongation, not static activator binding, triggers its writing. Third, MLL1 knockdown depleted H3K4me3 without compromising TNF-α or IL-1β induced transcription, showing that loss of H3K4me3 does not limit or impair transcriptional output. Together, these findings demonstrate that H3K4me3 operates as a post-transcriptional record of past synthesis rather than an instructive driver.
Using an inducible gene system, this study provides the first report in human gene models that H3K4me3 accumulation requires and strictly follows active transcription. While similar temporal lag between transcription and H3K4me3 deposition have been noted in non-human systems-such as yeast metabolic cycling and mouse liver circadian rhythms-our transcriptional perturbations and MLL1 knockdown experiments establish the precise functional sequence during acute human gene induction. By demonstrating that H3K4me3 accumulation occurs downstream of transcription, these findings reaffirm H3K4me3 as a post-transcriptional event rather than an instructive trigger. This framework offers a cohesive mechanism to address a long-standing paradox in chromatin biology: why acute depletion or catalytic knockout of H3K4me3 methyltransferases consistently fails to block initial gene activation or primary Pol II recruitment. Viewing H3K4me3 as a post-transcriptional consequence rather than a prerequisite trigger offers a plausible framework to help reconcile prior conflicting observations across eukaryotic systems.
A fundamental question arising from our study is whether delayed H3K4me3 loading is restricted to acute, inducible genes or operates similarly at constitutively active loci. Our kinetic dissection relied on the sharp “off-on-off” transcriptional burst of inflammatory loci, providing a clear window to uncouple transcription from histone modification. In contrast, assessing these kinetics at constitutively expressed genes presents a methodological challenge, as continuous transcription and delayed H3K4me3 exist in steady state rather than a resolvable pulse. While our transcriptional shutdown experiment at the constitutively active MYC locus sought to test whether delayed H3K4me3 deposition persists post-inhibition, the sustained mark presence observed after arrest could suggest this phenomenon. However, this persistence could reflect either continued active deposition or merely slow mark turnover and delayed clearance. Ultimately, determining whether post-transcriptional H3K4me3 accumulation is a universal principle across steady-state genomic regions will require future approaches combining metabolic pulse-chase histone labeling with rapid catalytic inhibition of methyltransferases or transcription.
If H3K4me3 does not drive primary initiation, what is its real post-transcriptional role? While our observation of delayed H3K4me3 loading does not definitively prove a singular downstream function, it raises compelling hypotheses regarding how this mark operates after transcription has commenced. Based on existing literature and our kinetic observations, we speculate that post-transcriptional H3K4me3 may contribute to several non-mutually exclusive processes: (i) preserving an open promoter architecture around the TSS to prevent the encroachment of repressive marks for chromatin barrier maintenance; (ii) genome integrity protection, by safeguarding exposed, open DNA at highly active promoters against transcription-induced damage; (iii) transcriptional memory, by serving as a molecular record of recent synthesis to “prime” the locus for more efficient reactivation during subsequent stimulation; and (iv) temporal reader recruitment, by acting as a late beacon to recruit nuclear reader proteins that coordinate splicing, elongation efficiency, or chromatin resetting upon termination.
Overall, our findings in human inducible gene models indicate that H3K4me3 deposition is temporally uncoupled from initial transcription at TNF-α and IL-1β, pointing toward a post-activation role rather than an instructive driver of initiation. These results provide a strong rationale for future studies to systematically evaluate whether similar kinetic uncoupling operates across broader genomic contexts and to define its precise post-transcriptional functions. Countless disease-profiling studies-spanning cancer, autoimmune conditions, and neurodegenerative disorders-have documented extensive H3K4me3 dysregulation. While our findings stem from acute response loci, this delayed-loading perspective offers a nuanced lens through which existing disease datasets can be re-interpreted. Applying this kinetic framework to disease-associated H3K4me3 patterns may offer valuable insights into disease mechanisms, gene aberrations, and chromatin locus resetting.
Ultimately, H3K4me3 represents just one mark within a vast landscape of PTMs, whose functional roles in diverse nuclear mechanisms remain incompletely understood. While comprehensive mapping initiatives have catalogued these marks across the genome, systematically decoding how PTMs dynamically coordinate these multi-step nuclear processes represents a critical frontier for epigenetic research and translational applications. However, translating these fundamental insights into targeted epigenetic therapies is currently hindered by the limited genome-wide specificity of conventional inhibitors, which can cause off-target toxicity and drug resistance. Overcoming these clinical barriers will require developing highly selective small-molecule chemical inhibitors targeting specific classes or domains of epigenetic modulators, as well as deploying locus-specific precision tools and cell-targeted formulations to restrict chromatin engineering strictly to disease-driving loci.